

The FDA has released draft guidance for the gene therapy community — and the window to shape it closes September 1, 2026.
The new Draft Guidance is titled: Leveraging Prior Knowledge in the Development of Human Gene Therapy Products Incorporating Genome Editing.
This is necessary reading for anyone in a development role at an organization that is pursuing human gene therapy products. This guidance is meant to assist organizations in speeding up development of products when prior knowledge is meaningful across the organization’s platform. The recommendations are focused on Gene Therapy but may be applicable to all Cell and Gene Therapies (CGT).
Accelerating development is critical as many of the CGT products are designed for unmet needs for rare diseases which commonly have smaller patient populations.
The guidance provides key definitions and a non-exhaustive list of recommendations for the audience.
The document is broken into sections discussing:
A) CMC
B) Non-clinical
C) Clinical
Each of these sections provides meaningful and targeted guidance for the respective areas. For example, the CMC section offers the example that prior knowledge “Prior knowledge may be considered for justifying which quality attributes to include in a lot release specification and suitable acceptance criteria for those quality attributes.” Lot specifications and Certificate of Analysis Testing are areas where requirements can rapidly balloon and lead to testing that does not actually provide value to the process or product. This is especially critical for GT products where the timelines for release are by necessity shorter than a more traditional small or large molecule pharmaceutical.
Process characterization is another potential area to use prior knowledge. Characterization takes a lot of time, product, and resources to execute. While patient populations tend to be small, the availability of products from the manufacturing process is also very small. Leveraging characterization data from another product will help organizations to move more products forward with the goal of discovering and developing innovative new products to treat these vulnerable patient populations.
The Clinical section provides the audience with general information about leveraging prior knowledge for trial design and trial conduct or analysis. Prior trial outcomes can be meaningful to improve design for future trials, but organizations should be interacting with regulators throughout development through filing to ensure that the approach is acceptable. Discovering that the approach to leveraging prior data was unacceptable to regulators very late in the process or at the time of filing would lead to significant rework and delays.
The final section of the document gives the reader details on what to submit when filing using leveraged prior knowledge.
This document would be of benefit for individuals who are responsible for developing, have oversight for clinical and non-clinical activities, or are regulatory professionals within the Gene Therapy spaces.
What Most People Are Missing
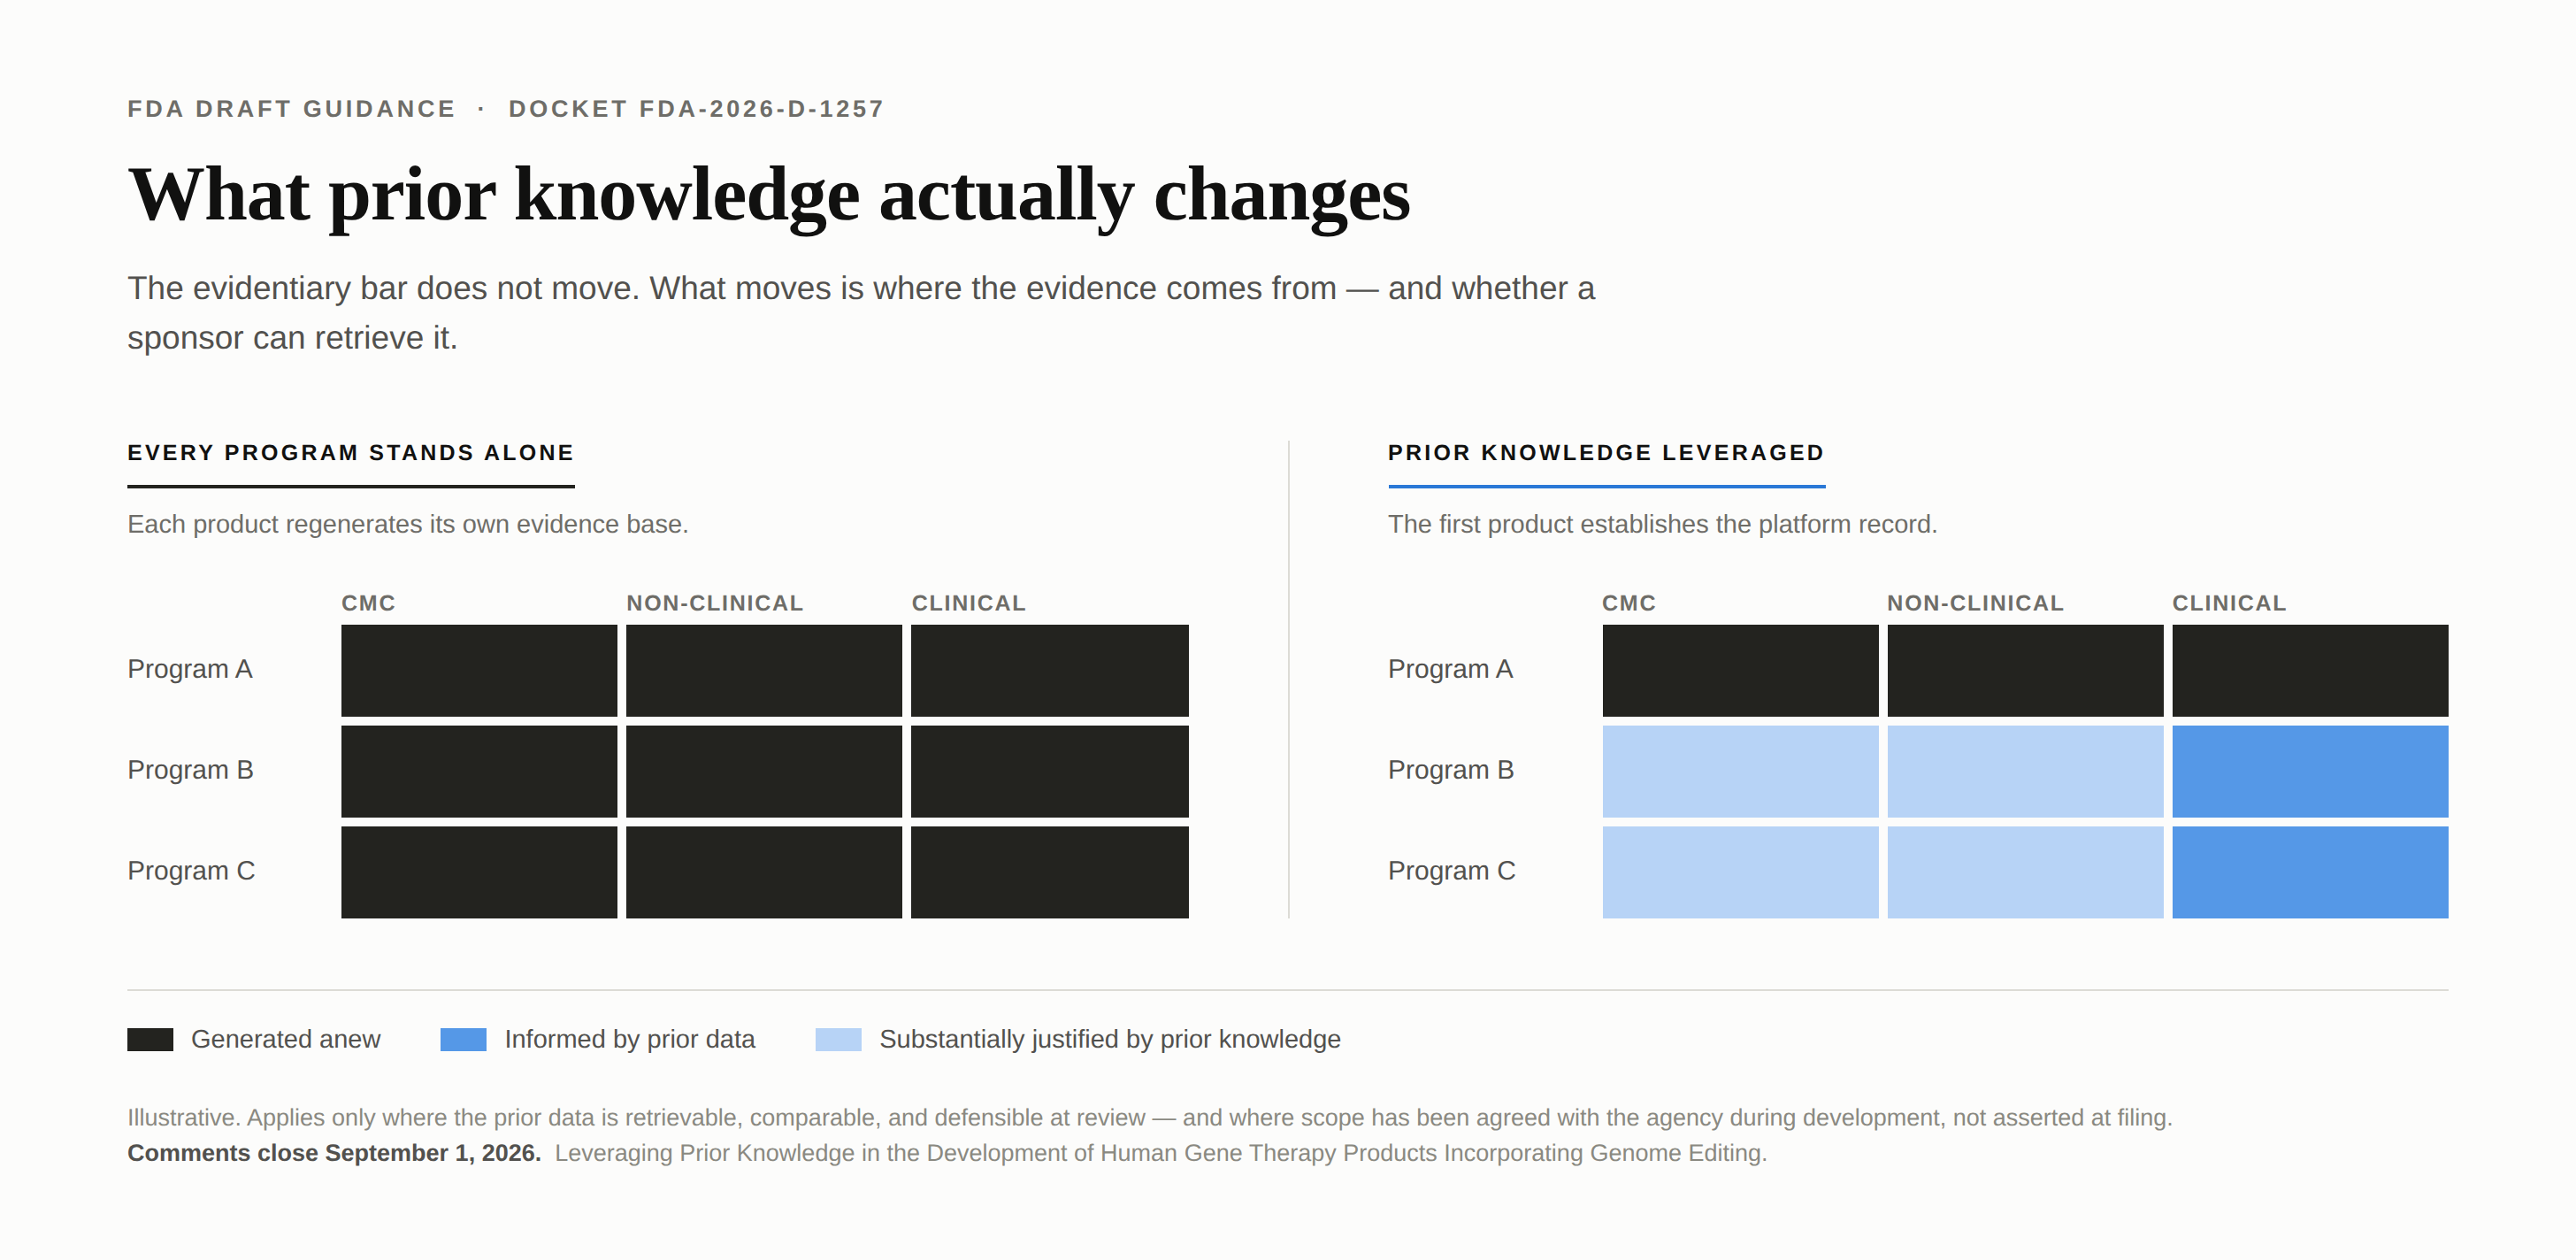
This guidance is being read as a regulatory concession — the FDA making life easier for gene therapy developers. That reading is incomplete. The agency is not lowering the evidentiary bar; it is allowing sponsors to meet that bar with data they have already generated. The burden does not disappear. It shifts — from generating evidence to retrieving, contextualizing, and defending it.
That shift is where most organizations will struggle, and it has almost nothing to do with regulatory strategy. Leveraging prior knowledge across a platform requires an organization to actually know what it knows. In practice, characterization data lives in program-specific silos: study reports in one system, raw analytical data in another, the scientific rationale for a critical decision in the head of a scientist who left two years ago. Reconstructing a defensible evidentiary thread across three programs and five years is a knowledge management problem, and knowledge management has been chronically underfunded in this sector relative to the value it is about to deliver.
The second-order effect is competitive. Two companies with equally strong science will not get equal benefit from this guidance. The one that invested in structured data, harmonized analytical methods across programs, and disciplined documentation will compress development timelines meaningfully. The one running on tribal knowledge will file substantially the same package it always would have — not because the pathway was unavailable, but because assembling the justification would cost more than repeating the study. Over two or three product cycles, that gap compounds into a durable advantage that has nothing to do with the underlying biology.
There is a corresponding failure mode worth naming. The temptation will be to claim prior knowledge broadly without the evidentiary spine to support it, and to discover at filing that the linkage between the prior product and the current one does not hold up under review. The guidance’s repeated emphasis on early and sustained interaction with regulators is the tell: the agency expects the scope of leveraged data to be negotiated prospectively, not asserted retrospectively. Sponsors who treat prior knowledge as something to be justified in the submission rather than agreed to during development are setting up exactly the late-stage rework this guidance is meant to prevent.
One open question deserves attention from anyone on the CDMO side. Prior knowledge is easiest to defend when it is your own, but a CDMO running genome editing processes across many sponsors accumulates platform experience no single client can match. Whether — and how — that pooled knowledge can be leveraged is not settled here, and the commercial agreements governing it were largely written before the question existed. Expect this to become a live negotiation point in CGT manufacturing contracts well before it becomes a regulatory one.
Comments can be submitted here. If your organization intends to weigh in, that work needs to happen now — once the docket closes, the next opportunity to influence this thinking is the final guidance, and by then the positions are set.
The document can be found on the US FDA Website.
Key Takeaways:
1. The FDA is establishing guidance on usage of prior knowledge for CGT processes and products.
2. Professionals in the Gene Therapy space are encouraged to review the draft document to understand the current thinking by the FDA on the leveraging of prior knowledge.
3. The community has only until September 1, 2026, to influence the final guidance — the comment window closes that day.
4. The practical constraint on using prior knowledge is not regulatory permission — it is whether your organization can retrieve and defend its own historical data. Assess that capability before you plan around it.
